Riepilogo del workflow di DE per RNA-Seq
NOTA: Il caricamento di questo esercizio potrebbe richiedere un po' più di tempo.
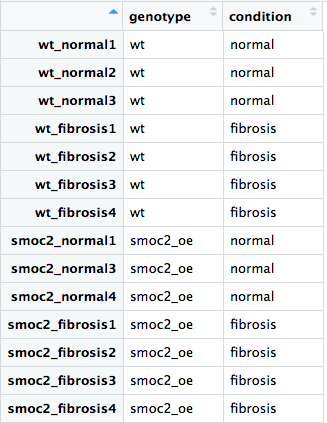
Ripercorriamo il workflow di DESeq2 usando l'intero dataset, che include sia i campioni wildtype sia quelli con overespressione di smoc2. Abbiamo già caricato per te le librerie DESeq2 e dplyr, oltre al file di metadati all_metadata e al file di conteggi grezzi all_rawcounts.
Questo esercizio fa parte del corso
RNA-Seq con Bioconductor in R
Istruzioni dell'esercizio
- Verifica che i campioni siano nello stesso ordine in
all_rawcountseall_metadatausandorownames(),colnames(),all()e l'operatore%in%. - Crea l'oggetto DESeq2 usando il design appropriato, testando l'effetto di
conditionmentre controlli pergenotype. - Crea l'oggetto DESeq2 usando il design appropriato, controllando separatamente per
genotypeecondition, ma testa pergenotype:condition.
esercizio interattivo pratico
Prova questo esercizio completando questo codice di esempio.
# Check that all of the samples are in the same order in the metadata and count data
all(___(___) %in% ___(___))

# DESeq object to test for the effect of fibrosis regardless of genotype
dds_all <- DESeqDataSetFromMatrix(countData = ___,
                        colData = ___,
                        design = ___)

# DESeq object to test for the effect of genotype on the effect of fibrosis
dds_complex <- DESeqDataSetFromMatrix(countData = ___,
                                ___,
                                ___)